Category Archives: Projects

Posts with category ‘Projects’ will appear under
the Project item in the Menu bar.

Pilot waves in CASINO: from toy boxes to real atoms, molecules and crystals

Everything you were taught at university about the weirdness of quantum mechanics, from cats that are simultaneously dead and alive to reality waiting politely for an observer to look at it before deciding what to do, hangs on a single word. In 1926 Max Born suggested that the squared modulus of the wave function, |Ψ|², gives the probability of finding a particle at a given place when you make a measurement. In 1927, at the fifth Solvay conference in Brussels, Louis de Broglie pointed out that if you change the word finding to being, every paradox evaporates. Both entities exist at all times: a real wave field Ψ evolving smoothly without collapse according to the Schrödinger equation, and real particles at definite positions guided along continuous trajectories by the gradient of the wave’s phase. David Bohm rediscovered the full many-body theory in 1952, John Bell championed it for thirty years, and most physics departments carried on pretending it did not exist.

Once you take the trajectories seriously, however, a remarkable physical question opens up. Because the de Broglie–Bohm velocity field carries the probability density |Ψ|² along with the wave, an ensemble of systems distributed as |Ψ|² today will remain distributed as |Ψ|² forever, reproducing every statistical prediction of textbook quantum mechanics. In pilot-wave theory, therefore, the Born rule is not a fundamental law of nature written on stone tablets; it is a state of statistical equilibrium, quantum equilibrium, strictly analogous to Maxwell’s thermal distribution in a classical gas. As Antony Valentini showed in 1991 with his subquantum H-theorem, an ensemble started out of equilibrium (P ≠ |Ψ|²) is stirred by the complex phase gradient of the wave until, on any coarse-grained scale, it relaxes towards |Ψ|².

From toy boxes to real matter: why CASINO is here

CASINO’s involvement in this story began in 2010 when I wrote LOUIS, a trajectory code designed to test Valentini’s relaxation theorem for a single particle in a two-dimensional box. In our 2012 paper (Proc. R. Soc. A 468, 990), Nick Russell, Antony Valentini and I showed how the nodes of a multi-mode wave function act as microscopic quantum vortices, winding an out-of-equilibrium probability density into ever finer filaments until it matches the Born rule. LOUIS has shipped as a utility inside CASINO since 2013, and in today’s CASINO v3.2 release it gains a new manual and a full ensemble driver.

Yet for thirty-five years, a glaring gap remained in the foundations literature. Every numerical study of quantum relaxation had been performed in a toy model: a particle in a box, a harmonic oscillator, or a free scalar field. Nobody had ever followed an out-of-equilibrium ensemble in a real, correlated many-electron atom, molecule or crystal. The reason was simple: foundations theorists did not have correlated many-electron wave functions, and the quantum Monte Carlo community, who spend their lives building Slater–Jastrow and multideterminant wave functions within fractions of a millihartree of exact eigenstates, never used them to integrate real-time trajectories. At every step of a variational or diffusion Monte Carlo run, CASINO already computes the exact complex vector ∇i ln Ψ for every electron: its real part is the familiar DMC drift vector, and its imaginary part is the de Broglie–Bohm guidance velocity! We had the machinery sitting in the code all along.

In CASINO v3.2, that machinery is now unleashed. Setting RUNTYPE = pilot_eval turns CASINO into a stateless, multi-state wave-function evaluator that computes ln|Ψ|, the phase, the local energy, the local kinetic energy (which, for a real state away from its nodes, is the Bohmian quantum potential Q), and the logarithmic gradients for almost any wave function CASINO knows how to read. The companion driver pilotdrive performs subspace diagonalisation across a set of correlated VMC trial states and integrates the real-time trajectories of an ensemble using an adaptive fifth-order Dormand–Prince stepper, or, when a diffusion constant is switched on, integrates Edward Nelson’s 1966 real-time stochastic mechanics. Evaluating the wave functions scales gently as about N1.6, taking tens of microseconds per step in helium and a few milliseconds in a 64-electron crystal supercell.

What real electrons taught us: from helium to diamond

I’ve just completed the first study of de Broglie–Bohm and Nelson relaxation across a five-rung ladder of real correlated matter: the helium atom (2 electrons in superpositions of up to ten singlet states within 0.6 mHa of exact), the beryllium atom (4 electrons, 12 states), the water molecule (10 electrons, 6 states), the benzene molecule (42 electrons, 4 states), and a periodic diamond crystal (64 electrons in a 2×2×2 supercell, 5 states). Turning QMC trial wave functions into real-time pilot waves immediately uncovered three pieces of physics that no toy box could show:

  1. The dimensional cascade. From beryllium to diamond, the initial drop in the coarse-grained H-function accelerates dramatically as the dimension 3N of the configuration space grows. Within just 20 atomic units of time (about half a femtosecond), H̄ falls to a third of its initial value in beryllium, an eighth in water, and under a fiftieth in benzene and diamond, with the 64-electron diamond crystal reaching the statistical sampling floor of a Born ensemble by the second recorded step (5 a.u.).
  2. Spontaneous Loschmidt echoes on a phase torus. In a few-electron atom like helium, adding more states does not guarantee faster relaxation. Because bound atomic eigenstates are real, whenever the relative phases 2(En − E1)t of the superposed states pass near multiples of 2π on their torus, a condition predictable from the energy spectrum alone before running a single trajectory, the wave function becomes nearly real up to a global phase. At that “near-real instant”, the velocity field becomes odd in time, v(t* + τ) ≈ −v(t* − τ): every electron comes to a halt and retraces its own path backwards through six-dimensional space! In five-state helium, the ensemble relaxes steadily for 556 atomic units down to a tenth of its initial H̄, hits the near-real instant at t* = 555.9 a.u., and dutifully climbs back up its own mirrored decay curve for the next 170 a.u. Under Nelson’s stochastic dynamics, by contrast, the osmotic drift and Brownian noise are not odd under time reversal: the noise obliterates the Loschmidt echo, fills the diffuse Rydberg halo by diffusion, and relaxes water and benzene six to eight times faster than the deterministic flow.
  3. The continuity defect and the Real-Jastrow Theorem. Because QMC trial wave functions are not exact eigenstates, differentiating |Ψ|² along a Bohmian trajectory reveals a small continuity leak whose rate ε(R, t) is driven by the exact local-energy fluctuations EL(n)(R) − En that variational Monte Carlo minimises. Integrating that defect along each trajectory yields an exact pathwise weight eS that restores the Born distribution. Even better, because a common real Jastrow factor eJ has zero phase, it cancels out of Im ∇ ln Ψ for a single state: the Jastrow factor carves a 1-Hartree correlation hole around each electron in the Bohmian quantum potential Q while leaving a single state’s velocity field completely untouched!

The films: imaginary time, real time, and attosecond charge migration

Having exact trajectories and walker histories on disk also allowed us to build a new rendering pipeline linking CASINO and Blender. When watching these animations, one fundamental distinction should be kept in mind.

The films of diffusion Monte Carlo (DMC), among them those of caffeine, C60 and small molecules whose nuclei are quantum particles, are on the Videos and Simulations page, with interactive 3D browser scenes. In those the motion is a computational random walk in imaginary time τ towards the ground state; in real time, the electrons of a real non-degenerate ground state stand completely still where the repulsive quantum force −∇Q balances the electrostatic force −∇V.

The five films below cross over into real time, showing what electrons actually do when guided by Nelson’s stochastic mechanics and by the deterministic de Broglie–Bohm pilot wave. Turn the sound up: every one of them has sound but the film of beryllium.

Full screen: the button among the film’s controls, at its lower right in most browsers, or in Firefox, Chrome and Edge a double-click on the film; on a phone, tap the film, then that button, and turn the phone sideways. Or open the film by itself, alone in the browser’s window.

Attosecond charge migration in chloroacetylene: 15 outer electrons shuttling a hole between a chlorine atom and a carbon triple bond every 1.2 femtoseconds, after one electron is taken suddenly from the orbital that lies mostly on the chlorine of H–C≡C–Cl. The nuclei are held still and the ion kept to the two of its states that make up most of what the removal leaves, whose beat moves the hole: amber where the ion has less electron density than the neutral molecule, blue where it has more. The streaks are the paths of those electrons, each moved by the de Broglie–Bohm guidance law with its spin a fixed label, integrated by CASINO’s pilotdrive, first in one molecule and then in 296 drawn together, whose electrons sample the ion’s density at the start and at every beat, the approximations letting the sample drift from it between beats; a second of film is about 167 attoseconds. Chlorine’s spin-orbit coupling is left out, and in the real molecule the nuclei move and the beat is expected to fade within a few femtoseconds.

Full screen: the button among the film’s controls, at its lower right in most browsers, or in Firefox, Chrome and Edge a double-click on the film; on a phone, tap the film, then that button, and turn the phone sideways. Or open the film by itself, alone in the browser’s window.

Edward Nelson’s stochastic mechanics (1966): in a real stationary state, where a de Broglie–Bohm electron sits still, a Nelson electron undergoes frictionless Brownian diffusion driven by the osmotic velocity Re ∇ ln Ψ—the exact same Markov process as the drift-diffusion step of importance-sampled DMC. Shown here for a single hydrogen 1s electron, an ensemble of 2,000 hydrogen atoms, and the 74 valence electrons of caffeine.

Full screen: the button among the film’s controls, at its lower right in most browsers, or in Firefox, Chrome and Edge a double-click on the film; on a phone, tap the film, then that button, and turn the phone sideways. Or open the film by itself, alone in the browser’s window.

Quantum relaxation in the beryllium atom: four electrons in a superposition of twelve singlet states, guided by CASINO’s correlated pilot wave. On the left, 4,000 atoms started out of equilibrium in the ground-state density; on the right, 2,000 atoms started in the Born distribution; beneath, the coarse-grained H-function falling over 200 atomic units (4.8 femtoseconds) with its characteristic beat pause between 70 and 115 a.u. The film has no sound.

Full screen: the button among the film’s controls, at its lower right in most browsers, or in Firefox, Chrome and Edge a double-click on the film; on a phone, tap the film, then that button, and turn the phone sideways. Or open the film by itself, alone in the browser’s window.

Vortex stirring in 2D (LOUIS): the classic 2012 calculation rendered afresh in high definition. An out-of-equilibrium ensemble of particles in a square box is guided by a 16-mode superposition; the moving nodes act as quantum tornadoes that stretch and fold the probability density into fine filaments until the Born rule is reached.

Full screen: the button among the film’s controls, at its lower right in most browsers, or in Firefox, Chrome and Edge a double-click on the film; on a phone, tap the film, then that button, and turn the phone sideways. Or open the film by itself, alone in the browser’s window.

A particle in a cubic box is guided by a wave made of sixty-four of the box’s standing waves with random phases, and an ensemble of such particles, started away from the wave’s intensity, is stirred to equilibrium by the wave’s nodes, which in three dimensions are lines that move, stretch and reconnect.

Where to find the code and the details

The kinship between imaginary-time DMC walks and real-time pilot-wave and Nelson trajectories is not merely a curiosity. Analysing the stochastic behaviour of walkers near nodal surfaces has already led in CASINO v3.2 to an exact three-dimensional Bessel-process near-node propagator for DMC. You can explore more films and interactive browser visualisations on the Videos and Simulations page, the background lectures on the de Broglie–Bohm page, the technical specifications of pilot_eval and pilotdrive in Chapter 16 (“The wave function outside QMC”) and Chapter 20 of the new CASINO v3.2 manual, and the commit-by-commit history in the development log.
Binding energies and electronic properties of two-dimensional materials

elahehneil

Elaheh Mostaani, Neil Drummond, Viktor Zolyomi and Vladimir Fal’ko

Two-dimensional (2D) materials are currently of great interest to the scientific community due to their extraordinary electronic and mechanical properties. Researchers are interested not only in monolayers of these materials, but also in few-layer and stacked systems, such as graphene on boron nitride. To model these systems it is necessary to understand the binding properties of layered materials. Unfortunately, standard first-principles methods such as density functional theory (DFT) are unable to describe the van der Waals interactions between 2D layers in a quantitative fashion. Instead we are using quantum Monte Carlo (QMC) methods to study the binding behaviour of 2D materials.

We have calculated the binding energy of bilayer graphene at the equilibrium separation, in both the AA- and AB-stacked arrangements. Our data will assist the benchmarking of van der Waals DFT functionals. We are now calculating the full binding-energy curve of bilayer graphene to provide a more complete description of the interaction of graphene sheets. We are also calculating the binding energy of graphene on boron nitride in various configurations, to support experimental work on graphene superlattices.

In addition, we are using QMC methods to study the electronic band structures of monolayer 2D materials such as boron nitride, transition-metal dichalcogenides and indium chalcogenides. Unlike DFT, QMC  does not suffer from underestimation of electronic band gaps. Furthermore, QMC can be used to calculate exciton binding energies, which are very significant in these 2D semiconductors. Our results will facilitate the characterisation of monolayers of these materials.

New benchmark paper for sI methane hydrate

Benchmarking the performance of density functional theory and point charge force fields in their description of sI methane hydrate against diffusion Monte Carlo

Stephen Cox, Mike Towler, Angelos Michaelides, Dario Alfè

clathrate

Preprint available: http://arxiv.org/abs/1402.6874

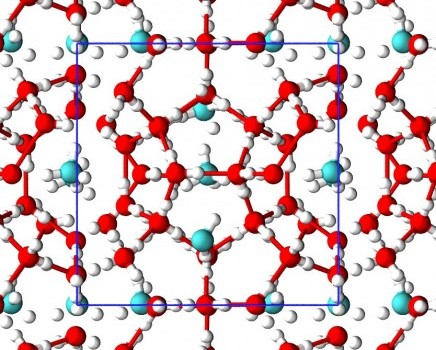
Due to the problems that gas hydrates pose to the energy industry (their formation blocks gas and oil extraction lines) and their potential as an untapped energy resource (there is an abundance of naturally occurring methane hydrate that exceeds conventional gas reserves by at least an order of magnitude), there has been much interest in modelling gas hydrates using computer simulation in recent years. To describe the interactions between the water and methane molecules present in gas hydrates, it is common to use approximations at the density functional theory (DFT) or force field (FF) levels. There is, however, a lack of high quality reference data to which the performance of these methods can be compared. This article remedies this by presenting high-quality diffusion Monte Carlo (DMC) data for bulk sI methane hydrate, the first such data for this type of system.

We also test a variety of commonly used DFT and FF methods against the DMC data. How to account for van der Waals dispersion forces in DFT is currently one of the hottest topics in computational chemistry and physics, and one of the main aspects of this work is investigating how dispersion corrected functionals affect the results relative to the local density and generalized gradient approximations. Although we find that dispersion corrected functionals improve some of the results, they do not guarantee an improved description of methane hydrate in all areas.

Not only will this work be of interest for developers of FFs and DFT, but also for anybody with an interest in using computer simulation to model methane hydrate that wishes to validate their methods.

Anharmonic vibrational properties of solids

165
Bartomeu Monserrat
, Neil Drummond, Richard Needs

The vibrational properties of solids are usually studied within the harmonic approximation, which is valid when the motion of the atomic nuclei is restricted to the neighbourhood of their equilibrium positions. This approximation is usually very good when investigating standard solids under normal conditions. However, for the lighter nuclei and at high temperatures, atoms explore regions far away from their equilibrium positions, and the harmonic approximation is no longer valid.

We have worked on a methodology to study the vibrational properties of solids beyond the harmonic approximation. We map the Born-Oppenheimer energy surface on which the atoms move along phonon modes, and then solve the resulting equations self-consistently within a mean-field formalism. We then calculate phonon expectation values from the anharmonic phonon wave function. This allows us to investigate physical properties such as the electronic band gap renormalisation due to electron-phonon coupling, or thermal expansion using the stress tensor [PRB 87, 144302 (2013)].

The first applications of this methodology have included the study of systems comprising of the lightest elements: hydrogen and helium. For hydrogen we have investigated the dissociation of molecular solid hydrogen at high pressure with quantum Monte Carlo calculations for the electronic part, and anharmonic vibrational calculations for the nuclear part. We have predicted a dissociation pressure of 370 GPa, within reach of static diamond anvil cell experiments in the near future. This work has been done in collaboration with Sam Azadi and Matthew Foulkes. For helium, we have calculated the effects of electron-phonon coupling on the metallization transition, predicting a strong zero-point correction and temperature dependence, both increasing the static lattice metallization pressure. This result has implications in the area of white dwarf cooling. This work has been done in collaboration with Chris Pickard.

Discovering new crystalline forms of metal oxides

lyle

Matthew J. Lyle, Chris J. Pickard, and Richard J. Needs

High performance computing is an essential part of many areas of scientific research. Materials discovery, in particular, is especially suited to computational simulation. Here we use an ab-initio random structure searching (AIRSS) algorithm to generate thousands of possible atomic arrangements for our system of choice. Constraints such as symmetry and stoichiometry can be incorporated to help our search towards particular structures we may be after. These candidate structures are energetically ranked, usually using density functional theory (DFT), and low-energy or other interesting structures are further investigated.

My projects have focussed on finding new crystalline forms of metal oxides, including alumina (Al2O3) and titania (TiO2). Alumina exists in a number of distinct crystalline forms that are widely used in industry; annual global production is 100 Mtonne, valued at approximately U.S.$32 billion. Conventional crystalline aluminas exhibit negligible porosity, though many of their industrial applications require high porosities and surface areas. This is usually achieved through the fashioning of mesoporous channels, though often at the expense of crystallinity which is known to increase alumina stability. Moreover, crystalline microporous silicates and aluminosilicates, e.g. zeolites, have long been used as adsorbents, ion exchangers, and catalysts.

image

In this project we performed an extensive first-principles computational search for new crystalline forms of alumina that are stable at ambient pressure. We found 147 unique structures with energies intermediate between those of the conventional alumina phases and identified a new class of crystalline aluminas which are up to 43% less dense. We attribute these low densities to significant amounts (up to 100%) of fivefold Al coordination and the formation of highly microporous zeolite-like channels. These are the first crystalline aluminas exhibiting such extensive fivefold Al coordination and surface areas, suggesting a new paradigm for the processing of alumina for industrial use.


MAGE: Materials Automated Generation and Exploration

107
Gareth Conduit

Through the stone, bronze, and iron ages the discovery of new materials has chronicled human history. The coming of each age was sparked by a chance discovery of a new material. Despite the central importance of materials in enabling new technologies, even today the only way to develop new materials is through experiment driven trial and error. We have developed the first tool [Patent GB1302743.8], MAGE, Materials Automated Generation and Exploration: computationally designs materials with specified physical properties. We have proven its accuracy by predicting five new alloys [Patent GB1307533.8, Patent GB1307535.3, Rolls-Royce Group plc invention submissions NC12261, NC13006 & Acta Materialia, 61, 3378 (2013)] that were subsequently experimentally verified. MAGE can help not only materials scientists, but moreover design engineers. At present engineers must design new objects and products around the shortcomings of pre-existing but non-ideal materials. The capability to develop materials computationally would allow engineers to instantly optimize bespoke materials for their application, bringing materials into the heart of the design process.

The alloys developed include a new nickel-based alloy for turbine discs in jet engines, a novel alloy for lining the combustion chamber of a jet engine, and a molybdenum based forging alloy. Each alloy has eight individual physical properties that are predicted to match or exceed commercially available alternatives including fracture toughness, oxidation resistance, yield strength, creep resistance and processibility. These properties are calculated using a variety of techniques including ab initio calculations, physically based models, and interpolation of existing experimental data. Several properties for each alloy have subsequently been experimentally verified so the alloys are now undergoing compliance testing by Rolls Royce plc.

In the future we plan to further incorporate first principles methods into MAGE. Working with Samsung Electronics Co Ltd through a Global Research Outreach Grant we will first focus on designing new GaInN-based LEDs materials, exploiting Density Functional Theory to predict properties from first principles. This investigation should allow us to not only design potentially important semiconductor materials, but moreover act as a launch-pad to guide the future design of new materials from first principles.

A Stochastic Approach to Battery Design

ajm1
Martin Mayo, Hugh Glass, Andrew J. Morris.

LIBs are the rechargeable battery of choice for electric vehicles and portable electronic devices. There is substantial interest in enhancing the capacity of LIBs, driven by the economic and environmental advantages of increasing the range of electric vehicles, and enabling longer-life portable electronic devices. The traditional anode is composed of graphite but recently silicon has been proposed as an alternative which has a theoretical capacity some ten-times larger. Silicon anodes are not fully understood however and suffer from large volume expansion on charge and hysteresis on charge cycling. Phosphorous is another such candidate for high-capacity anodes due to its multiple allotropes (white, red and black phosphorus), its ability to make Hittorf-chains and novel structures when mixed with lithium.[1]

P21/c Li1Si1

The creation of new materials is both difficult and expensive. It is very difficult to “see” the structure of these materials over the length scales that they work and very expensive to create prototype materials to test. Whilst experimental physics can use x-rays, high-energy electrons or neutrons to infer the structure of these materials, this inference is made LiP double helix infinite chainmuch more robust when combined with theoretical predictions of the kinds of structures that can be formed. In a computer we use quantum mechanical calculations to simulate the results of these kinds of experiments, helping to understand materials and suggest new materials with the kind of properties desired. The theoretical prediction of even very simple structures has, until recently, been out of bounds due to the large number of possible atomic arrangements. The ab initio random structure searching method (AIRSS) [2] uses a stochastic approach to suggest different structural configurations of atoms within a material. By searching over a range of stoichiometries it is possible to model how a battery is charged as we can predict the structural changes of the electrodes as a charging potential is applied. Using this knowledge it is possible to perform further theoretical analysis such as NMR and EELS (electron energy loss) spectroscopies and predict charge and discharge voltages.

[1] “Inorganic Double Helix Structures of Unusually Simple Li-P Species”, Alexander S. Ivanov, Andrew J. Morris, Konstantin V. Bozhenko, Chris J. Pickard and Alexander I. Boldyrev, [Angewandte Chemie 51 33, 8330-8333 (2012)]

[2] C.J. Pickard, and R.J. Needs , Phys. Rev. Lett. 97, 045504 (2006)

Quantum Monte Carlo simulations of solid hydrogen

120

Jonathan Lloyd-Williams, Bartomeu Monserrat, Pablo López Ríos, Neil Drummond, and Richard Needs

Hydrogen has the simplest electronic structure of any atom but its bulk properties are surprisingly complex. As hydrogen can only form a single covalent bond, it is expected to remain molecular to very high pressures. Several solid phases of hydrogen have been observed; the low-pressure phase I which consists of a quantum crystal of rotating molecules on a hexagonal close-packed lattice transforms at pressures of about 110 GPa to the broken symmetry phase II, in which the mean molecular orientations are ordered, and then to phase III at about 150 GPa. A new phase IV was recently observed in room temperature experiments and has generated great excitement within the field. Despite many years of intensive study, the arrangements of the molecules in phases II and III remain undetermined. The structure of phase IV is also unknown.

Experimental measurements have only been able to provide limited information about the molecular orientations in phases II and III of hydrogen. The information obtained from X-ray diffraction is limited because of the weak scattering of hydrogen atoms. So far the most important experimental data have come from infra-red and Raman vibrational spectroscopy. Theoretical identification of the structures is also difficult because of the need to consider the quantum statistics of the nuclei, the large zero-point vibrational energies, and the small enthalpy differences between competing phases.

To make progress in understanding the high pressure phase diagram of hydrogen, we will perform diffusion Monte Carlo (DMC) calculations for the candidate phases (P21/c-24, C2/c-24, Cmca-12, Cmca-4, and Pc-48) at several different densities which will allow us to evaluate the enthalpy. DMC is a zero-temperature method but estimating the small room temperature corrections should be straightforward. We will first perform a set of static lattice DMC calculations and add zero-point motion effects from density functional theory to allow some preliminary comparisons with experiment. We will then perform a second set of calculations including the full zero-point motion effects within DMC.

Project begun: January 2013

Expected completion: December 2014

A pseudopotential for ultracold atomic gases

Pascal Bugnion, Gareth Conduit, Richard Needs

Ultracold atomic gases provide a test bed for the study of fundamental quantum condensed matter phenomena. We are particularly interested in fermionic cold atom gases as these can be used to model collective electronic processes in the solid state.

The dynamics of a dilute ultracold atomic gas is dominated by short-range interactions between the atoms, typically characterized by the scattering length of the inter-particle potential. Modelling this regime is challenging for QMC methods as the wave function diverges when particles coalesce.

Previous attempts at circumventing this divergence have normally involved using an effective potential to model the inter-particle interactions. This effective potential is constructed to reproduce some of the features of the true potential while avoiding its pathological behaviour. We investigate the applicability of the wide range of methods developed by the electronic structure and chemistry community to the development of an effective potential for interactions between cold atoms.

Pairing wave functions for quantum Monte Carlo

Pascal Bugnion, Pablo López Ríos, Richard Needs

Diffusion Monte-Carlo (DMC) has been used to describe many real and model systems, usually displaying impressive agreement with experiments while maintaining favourable scaling with system size.

The accuracy of diffusion Monte-Carlo is ultimately limited by the so-called ‘fixed-node’ error, which depends on the quality of the trial wave function used as input to the algorithm.

Typically, the DMC trial wave function is generated in a variational quantum Monte Carlo (VMC) calculation, where it is optimized to minimize the VMC energy. It is heuristically believed that trial wave functions with lower VMC energies will have better nodal surfaces.

To reduce fixed-node error, we therefore need VMC trial wave functions whose functional form is close to the ground state wave function. In this project, we will study a class of such wave functions: pairing wave functions. Pairing wave functions contain parameters which can be optimized variationally, allowing in principle the determination of a better nodal surface. This produces lower DMC energies and more accurate results. We shall investigate how well pairing functions behave in larger systems, looking in particular at size extensivity.