CASINO is a program for quantum Monte Carlo calculations of the electronic structure of atoms, molecules, surfaces and solids. It computes the properties of interacting electrons by solving the many-electron Schrödinger equation with stochastic methods, working with the many-electron wave function itself, as a function of the coordinates of every electron, rather than through an approximate functional of the density. Its results carry a statistical error bar, which is estimated with them, and a short list of systematic errors that can be named, tested and reduced.
What it computes
Two methods do most of the work. Variational Monte Carlo evaluates the energy, and other properties, of an explicitly correlated trial wave function and optimises its parameters; its accuracy is that of the trial function. Diffusion Monte Carlo then improves on it, projecting out the lowest-energy state that shares the nodal surface of the trial function, by a random walk of a population of electron configurations in imaginary time. CASINO also implements reptation Monte Carlo, which samples pure rather than mixed expectation values.
What these methods compute best is the total energy, and from it energy differences: binding, cohesive, formation and excitation energies, reaction barriers and equations of state. Densities and spin densities, pair-correlation functions, structure factors, momentum densities, density matrices, dipole moments, the localisation tensor of a periodic system and, for molecules in Gaussian basis sets, the forces on the atoms can be computed too, at more cost or with biases of their own. The systematic errors of DMC are the fixed-node approximation, by which the nodes of the trial function are imposed on the answer; the finite time step; the finite population of walkers; finite-size effects in periodic systems; and, where they are used, pseudopotentials. The last four can be measured and extrapolated away; the fixed-node error can only be reduced, by a better trial function, and running quantum Monte Carlo well is largely a matter of attending to all five.
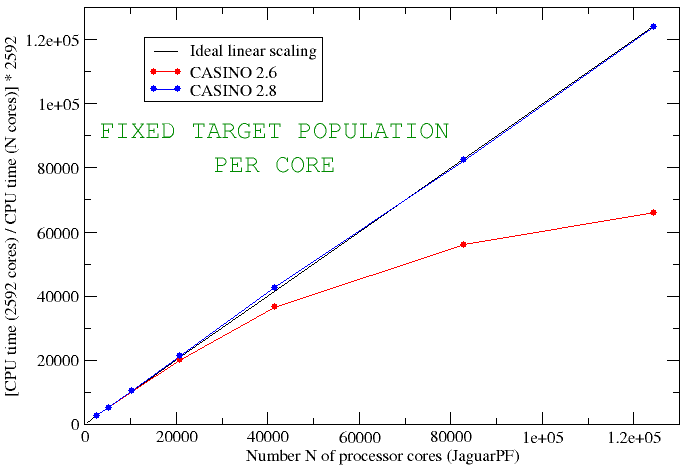
The method is expensive: the cost of a given error bar on the total energy rises as the third power of the number of electrons, the fourth with backflow, and steeply with atomic number in all-electron work. In exchange, the calculations parallelise almost perfectly, and CASINO has run efficiently on several hundred thousand processor cores.
What it is good for
Quantum Monte Carlo is most useful where electron correlation decides the answer and cheaper methods give a spread of answers depending on the approximation chosen: binding, cohesive and adsorption energies, the relative stability of structures and defects, weakly bound systems, and reference values against which faster methods can be tested. It also suits systems that other methods find awkward: the electron gas in one, two and three dimensions, whose quantum Monte Carlo energies, for the three-dimensional gas, are what the local density approximation of density functional theory was fitted to; electrons and holes in semiconductors; excitons and their complexes; positrons; and particles of arbitrary mass and charge.
What the program offers
CASINO treats finite systems and systems periodic in one, two or three dimensions, with real or complex wave functions, spin-polarised and, for some cases, non-collinear. Its trial wave functions are of Slater-Jastrow form, with optional multiple determinants, backflow and pairing functions, built from orbitals expanded in Gaussians, plane waves, blip functions, Slater-type functions or numerical grids. The orbitals come from other programs: in Gaussian basis sets from CRYSTAL, GAUSSIAN, GAMESS-US, MOLPRO, CFOUR, DALTON, NWCHEM, OpenMolcas, ORCA, PSI4, PySCF, Q-Chem and TURBOMOLE, and through TREXIO files from PySCF and Quantum Package; in plane waves from PWSCF (Quantum ESPRESSO), ABINIT, CASTEP, QBOX, MCEXX, GP/JEEP and K207; as Slater functions from ADF; and as numerical orbitals of atoms from ATSP2K and of diatomic molecules from 2DHF. The interfaces page describes each. Periodic Coulomb interactions are computed by Ewald sums or by the model periodic Coulomb interaction, which is faster and has smaller finite-size errors; non-local pseudopotentials are treated by the locality approximation or by Casula’s T-move scheme, and a library of pseudopotentials made for quantum Monte Carlo is provided.
Also there: optimisation of the trial wave function by variance or energy minimisation; compressed multideterminant expansions; twisted boundary conditions and twist averaging; the fixed-phase approximation for systems in a magnetic field; core-polarisation potentials; localised orbitals, with which the time to move every electron grows only in proportion to their number; and coupled DFT and DMC molecular dynamics by the Grossman-Mitas method. The manual’s first chapter lists every facility and how far each is developed.
The program is written in Fortran and runs in parallel with MPI, with OpenMP threads and shared memory as further options; it runs equally on a laptop and on the largest machines. It comes with a manual that explains the methods as well as the options, worked examples, a help system that describes every keyword, a set of test calculations with their reference results, and a large set of utilities for preparing input, running calculations and analysing their output.
Who made it
The research programme that led to CASINO was begun in the early 1990s by Richard Needs in the Theory of Condensed Matter group at the Cavendish Laboratory, Cambridge, with Guna Rajagopal, Matthew Foulkes and a number of students and postdoctoral researchers. Mike Towler, who joined the group late in 1996, gathered their test programs into a single program for atoms, molecules and periodic systems, and the first public version of CASINO appeared in 1999. Neil Drummond, from 2002, and Pablo López Ríos, from 2004, made fundamental improvements and extensions to it, and these four are its principal authors; many others have contributed over the years, and are named in the manual. The program is maintained by Mike Towler, with Neil Drummond and Pablo López Ríos.
CASINO 3.0 was released on 15 August 2026, the first release since 2013. Development has been continuous since: the beta offered at download is rebuilt each time a change is published, every change is described in the public DIARY, and the things-to-do page says what has been done and what is planned.
CASINO is free for academic and other non-profit use: request a copy, and please cite it in work that uses it. Questions go to the discussion forum, and the reading list is the place to start learning the method.